


CRISPR Genome Editing Solutions
VectorBuilder offers a comprehensive collection of CRISPR products and services for in vitro and in vivo genome editing. We provide ready-to-use CRISPR reagents including plasmid vectors, viruses, IVT RNAs and LNPs, pooled libraries, stable cell lines, and more for various experimental needs.
Highlights

Customizable: Our free, highly intuitive online vector design studio enables unlimited designs of your CRISPR vectors.

Comprehensive: Knockout, knockin, and CRISPRa/i with Cas9 variants available in various gene delivery formats.

Streamlined: Full service from experimental design through stable cell line engineering and library screening.

Expertise: Excellent quality, fast turnaround, and competitive pricing with powerful technical support.
What We Offer

CRISPR Vectors

CRISPR Virus

IVT RNA and LNP

Cas9 Protein

CRISPR Research Discovery Services
Empowered by our free, user-friendly online design platform, you can easily design and order custom and premade CRISPR vectors for all kinds of gene editing and regulation experiments.

All-in-one and dual-vector formats available in a variety of non-viral, viral, and transposon backbones for targeted knockout and knockin.

Optimized CRISPRa and CRISPRi systems for efficient endogenous gene activation or inhibition.

A rich collection of premade Cas9 vectors, gRNA vectors, and helper vectors for CRISPRa/i ready for immediate shipment as E. coli stocks.
VectorBuilder offers premium-quality virus packaging services for achieving highly efficient CRISPR targeting in difficult-to-transfect cells. Our proprietary technologies in virus packaging ensure improved titer, purity, viability, and consistency.

Scales ranging from 106 to 109 TU for research-grade packaging; 3rd generation lentivirus with various pseudotyping options available.

Scales ranging from 1010 to 1014 GC for research-grade packaging; more than 18 widely used serotypes available.

Scales ranging from 1010 to 1012 IFU; human Ad5, chimeric Ad5/F35, and gutless adenovirus available.
You can easily design gRNAs using our proprietary gRNA design tool to ensure efficient and specific gene targeting and order high-quality IVT gRNA and Cas9 mRNA for transfection or microinjection.

VectorBuilder offers Cas9 proteins optimized for direct transfection and high editing performance.

VectorBuilder offers custom and premade CRISPR libraries and stable cell lines for high-efficiency functional genomics.

We offer design, cloning, and packaging of pooled libraries as well as in vitro and in vivo functional screening and deconvolution.

Single and dual-gRNA lentivirus libraries for various screening purposes with whole genome and pathway scales in human and mouse.

Gene knockout, knockin, or point mutation using CRISPR with comprehensive QC.
A range of premade, functionally validated Cas9 expressing cell lines ready for functional study and high-throughput screening.

VectorBuilder is the one-stop provider for all your custom CRISPR needs.
Contact UsDesign Tips
| Category | Recommendation |
|---|---|
| CRISPR vector components |
Find more information here. |
| gRNA |
Find more information here. |
| Cas9 |
|
| PAM and gRNA compatibility |
|
| Reporter systems |
|
| Delivery systems |
Find more information here. |
| Donor DNA selection |
Find more information here. |
| Target site |
|
Technical Information
 CRISPR-mediated genome editing
The CRISPR system can be used for a growing variety of applications, and each CRISPR approach requires at least two basic components: a Cas protein and guide RNA (gRNA). The most commonly used Cas endonuclease is engineered from Streptococcus pyogenes (a.k.a. SpCas9 or, with codon optimization for expression in human cells, hCas9). It is an RNA-guided DNA nuclease which can generate double-strand breaks (DSBs) at target sites (Figure 1). Cas9 is localized to a target site in the host genome by the complementary gRNA, and the DSB is introduced if the gRNA was designed to target a region immediately adjacent to a Protospacer Adjacent Motif (PAM). The PAM sequence is dependent on the particular Cas enzyme being used: for SpCas9, the PAM sequence is NGG or NAG (orange DNA in Figure 1).
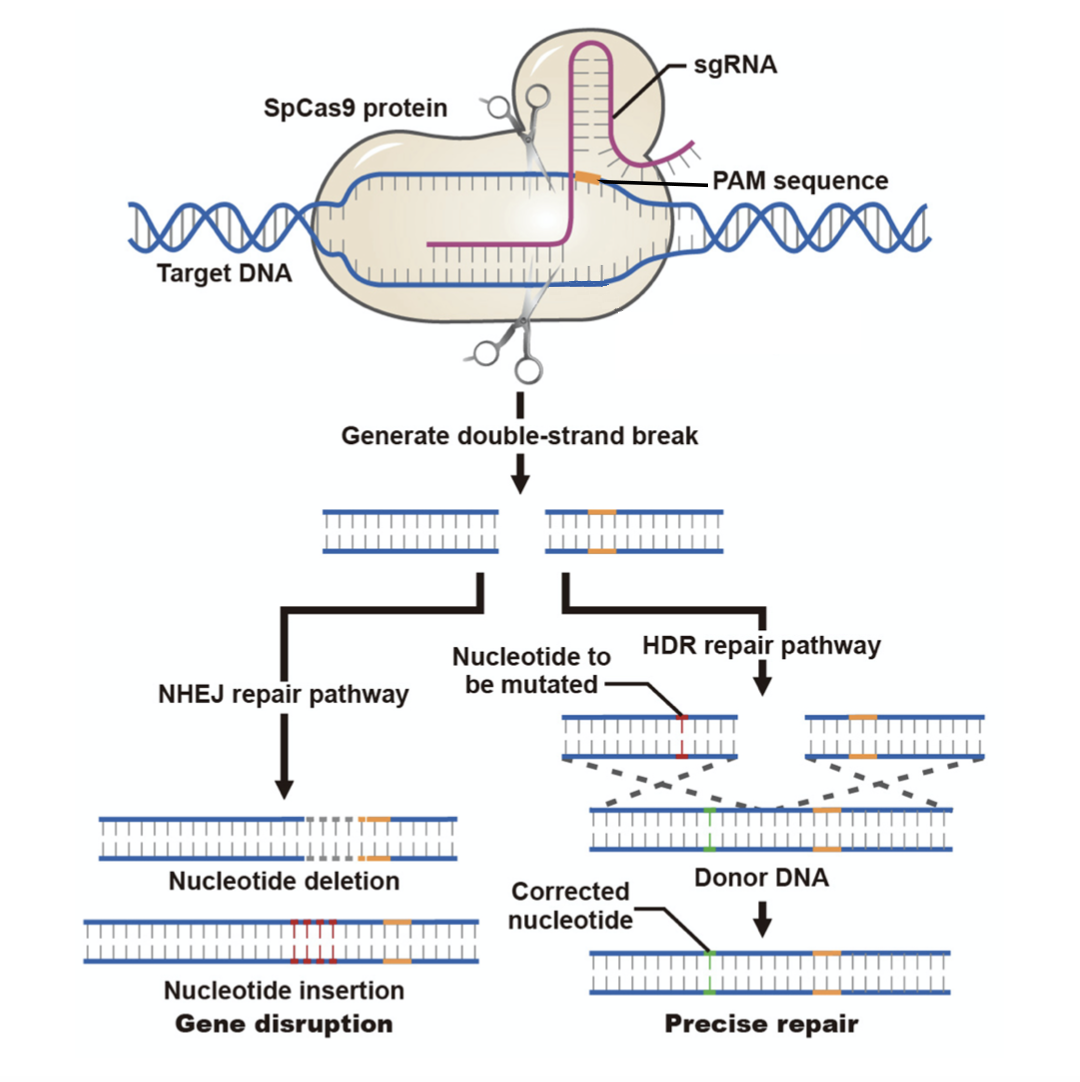
Figure 1. Mechanisms of CRISPR-induced DNA repair to produce gene knockout or precise sequence changes.
Once DSBs are generated by the CRISPR system, cells most commonly activate the non-homologous end joining (NHEJ) pathway to repair the DNA breaks, which usually results in small random deletions, or more rarely insertions and base substitutions. When these mutations disrupt a protein-coding region (e.g. a deletion causing a frameshift), they may lead to functional gene knockout. Dual gRNAs can be introduced with Cas9 to create two DSBs, thereby deleting a DNA fragment located between the two target sites (Figure 4), or dual gRNAs can be used to target two different genes simultaneously.
Alternatively, and less efficiently, DSBs can be repaired via the homology-directed repair (HDR) pathway when a DNA template is present. When donor DNA is introduced with the CRISPR components, HDR can result in replacement of the target genomic DNA sequence with the donor sequence, generating precise changes such as point mutations or knockin. The donor DNA template can be a single-stranded oligo nucleotide (ssODN) or a dsDNA fragment (usually linearized DNA derived from either regular plasmid or AAV). ssODNs are suitable for introducing point mutations or small tag insertions, while dsDNA fragments are widely used to introduce large fragment knockins like targeted stable integration of transgenes.
Another widely used Cas9 variant, Cas9 “nickase” (e.g. Cas9(D10A)), generates single-stranded cuts in DNA. If Cas9 nickase is used in conjunction with two gRNAs targeting the two opposite strands flanking a single target region, the nickase will generate two single-stranded cuts, one on each strand, resulting in wide-spread DSBs around the target region as shown in Figure 2. Because single nicks are generally repaired with high fidelity, efficient cleavage occurs only when both offset gRNAs bind correctly, creating a staggered DSB. This double-nicking strategy, therefore, dramatically reduces off-target activity while maintaining high on-target efficiency at many loci.

Figure 2. Nickase activity with two gRNAs.
 CRISPR-mediated gene regulation
A catalytically inactive form of Cas9 (dCas9) can be combined with different transcriptional complexes to achieve CRISPR-mediated transcriptional regulation of endogenous genomic loci. CRISPR-mediated gene regulation can be used for a wide variety of applications including testing regulatory elements on chromatin, reprogramming of target cells, or for whole genome screening to identify genes that drive proliferation and differentiation.
To activate gene transcription (CRISPRa), the Synergistic Activation Mediator (SAM) system can be used. This system utilizes three components: a dCas9/VP64 fusion protein, an MS2/P65/HSF1 helper complex, and a modified gRNA (msgRNA). The dCas9 protein is engineered with VP64, a synthetic transcriptional activator, and the gRNA is modified to include MS2 aptamers which recruit the MS2 complex consisting of the transactivation domains of NF-kB (p65) and HSF1. Together, the dCas9/VP64 and MS2/P65/HSF1 complexes work synergistically to recruit endogenous transcriptional machinery and cofactors to activate target gene expression, as shown in Figure 5. Beyond the SAM system, VectorBuilder offers the dCas9/VP64-p65-Rta (VPR) system which replaces the transactivation domain of HSF1 with the Epstein-Barr virus R transactivator. The dCas9/VPR system has been reported to exhibit higher levels of endogenous activation, although due to it's smaller size, dCas9/VP64 is preferable for AAV.
In contrast, to repress target gene transcription, dCas9 can be engineered with a transcriptional repressor, such as a Kruppel-associated box domain (KRAB) and MeCP2. Two versions of the dCas9/KRAB helper vector are currently available: the original dCas9/KRAB vector and an improved dCas9/KRAB/MeCP2 version which drives the expression of dCas9 fused to a bipartite repressor domain KRAB/MeCP2 for achieving more potent transcriptional repression of DNA target sites. The KRAB domain is commonly found in human transcription factors and is one of the most potent transcriptional repressors currently identified in the human genome. MeCP2 further enhances KRAB-mediated repression through recruitment of histone deacetylases, resulting in chromatin condensation and epigenetic-mediated repression. When the gRNA is designed to recruit dCas9/KRAB/MeCP2 to an endogenous promoter, as shown in Figure 6, transcription is heavily repressed.
 CRISPR delivery approaches
CRISPR-mediated genome editing or regulation requires target cells to co-express Cas9 and a target site-specific gRNA. Regardless of the application, CRISPR components must be introduced into target cells. Specific components can be transfected or transduced via different approaches including:
- gRNA and Cas9 plasmid
- gRNA and Cas9 virus (i.e. lentivirus, AAV, adenovirus, etc.)
- Mixture of gRNA and Cas9 mRNA
- Preformed RNP consisting of gRNA and Cas9 protein
Figure 3 summarizes the various methods for CRISPR delivery and the case studies in Figures 7, 8, 9 and 10 illustrate the effectiveness of each delivery method.
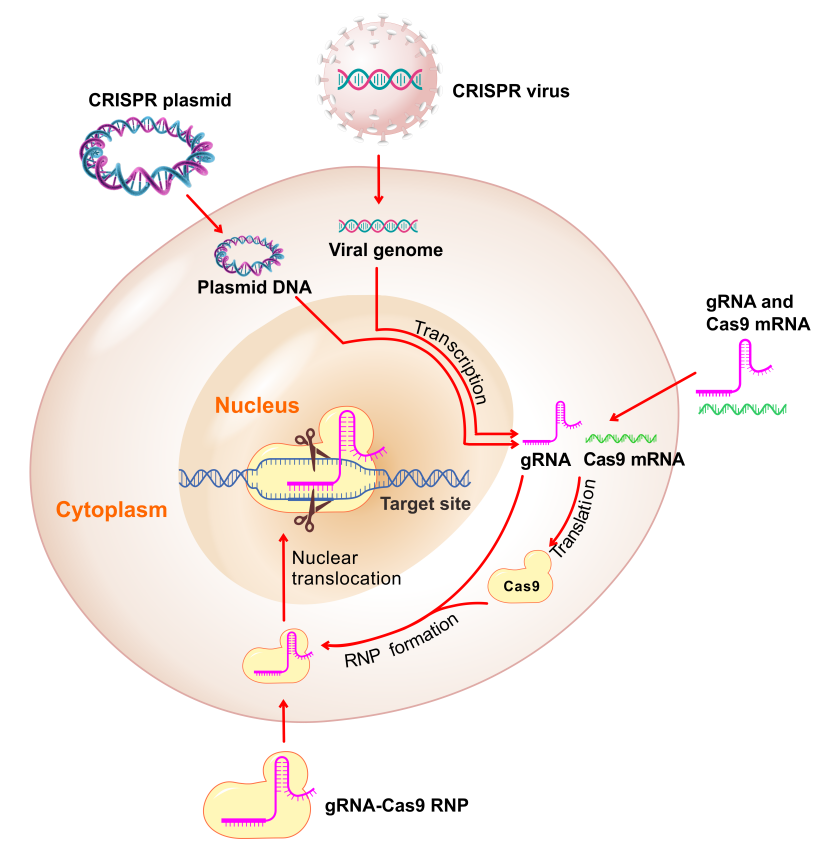
Figure 3. Popular methods for delivering CRISPR components into target cells.
The table below lists key advantages and disadvantages for each delivery approach and can be used as a reference to help you decide the most suitable approach for your experiments:
| Delivery Approach | Advantages | Disadvantages |
|---|---|---|
| gRNA and Cas9 plasmid |
|
|
| gRNA and Cas9 virus |
|
|
| Mixture of gRNA and Cas9 mRNA |
|
|
| Preformed gRNA-Cas9 RNP complex |
|
|
 gRNA databases
VectorBuilder’s online CRISPR vector design tool features optimized, whole-genome gRNA databases for human, mouse, and rat enabling you to design CRISPR vectors with high targeting efficiency. We are also expanding our gRNA database to include more Cas enzymes and more host species. We follow the algorithm utilized in CRISPR library design (CLD) to calculate specificity scores for gRNAs. Briefly, for a given gRNA intended to target a N(20)NGG sequence in a species, we search for all potential off-target sites in the genome of that species that have ≤3 mismatches with the target sequence. For each potential off-target site identified this way, a single off-target score is calculated. Scores for all the off-target sites are then used in aggregate to calculate the final specificity score of the gRNA, which is between 0 and 100, with higher values indicating greater targeting specificity. Please note that specificity scores are a theoretical guide. Actual targeting efficiency and specificity could depart from what the scores predict. gRNAs with low scores may still work well.
When you design CRISPR vectors on VectorBuilder’s online platform, you will have the option to search for your target genes in our database. Upon entering your gene name, you will see detailed information on all available gRNA designs against your target gene available in our database.
Case Studies
VectorBuilder offers a comprehensive and thoroughly validated range of CRISPR technologies, supporting diverse applications.
Several methods are available for delivering CRISPR components into target cells, including plasmid DNA, viral vectors, mRNA, and ribonucleoprotein (RNP) complexes, and our proven expertise across all of these platforms enables us to select and optimize the most effective approach for each application.
Dual-gRNA CRISPR gene knockout

Figure 4. Dual-gRNA CRISPR editing enables efficient gene knockout (KO). (A) Editing RNPs targeting separate sites on the GOI were electroporated into cells to delete a 13 kb region Single clones were then isolated and screened using PCR and Sanger sequencing. (B) Four primers, P1 to P4, were used in three PCRs to differentiate KO and WT clones. Clone 1 was validated as a homozygous KO mutant based on (C) PCR and (D) sequencing results.
Transcriptional activation with CRISPRa

Figure 5. Lentiviral CRISPRa-mediated upregulation of target gene expression. NIH3T3 cells stably expressing (A) SAM complex dCas9/VP64 and MS2/P65/HSF1 were transduced with lentivirus encoding for (B) msgRNA targeting the promoter region of the mouse Brn2 gene. (C) Relative gene expression of Brn2 in NIH3T3 cells transduced with either the targeting msgRNA, a scramble control, or no treatment control (NC) was measured by qRT-PCR (mean±SD, *P<0.05, ANOVA with Tukey’s post hoc test).
Transcriptional inhibition with CRISPRi

Figure 6. Lentiviral CRISPRi-mediated downregulation of target gene expression. Jurkat cells stably expressing the (A) dCas9/KRAB/MeCP2 transcriptional repressor complex were transduced with lentivirus encoding for (B) gRNA targeting the promoter region of the human CXCR4 gene. Following antibiotic selection, CXCR4 expression levels in Jurkat cells transduced with either the targeting gRNA, a scramble control, or no treatment control (NC) were measured by (C) Western blot and (D) qRT-PCR (mean±SD, ***P<0.001, ****P<0.0001, ANOVA with Tukey’s post hoc test). (E-F) Surface CXCR4 expression was quantified by flow cytometry, showing a ~50% reduction in cells transduced with the CXCR4-targeting gRNA compared to scramble control.
Non-viral delivery

Figure 7. Simplified gene editing with the all-in-one CRISPR system. (A) Regular plasmid expressing Cas9:T2A:mCherry and EGFP-targeting gRNA was transfected into HEK293T cells with stable EGFP expression (HEK293T-EGFP). EGFP expression was measured by flow cytometry 72 h post-transfection. (B) Representative fluorescence microscopy images, 100X. (C) Relative EGFP expression quantified by flow cytometry (mean±SD, ANOVA with Tukey’s post hoc test). (D) The targeted genomic region was PCR-amplified and editing was confirmed by T7E1 assay.
Viral delivery with lentivirus, AAV, or adenovirus

Figure 8. Stable and efficient gene editing with the all-in-one lentivirus-based CRISPR system. (A) Lentivirus expressing Cas9:T2A:Puro and EGFP-targeting gRNA was packaged and used to transduce stable EGFP-expressing HEK293T cells (MOI = 10, 50). (B) Representative fluorescence microscopy images, 100X. Efficiency of gene editing was measured using (C) T7E1 assay of PCR-amplified targeted genomic region and (D-E) flow cytometry, indicating a a ~70-90% reduction in EGFP expression depending on MOI used (mean±SD, ***P<0.001, ANOVA with Tukey’s post hoc test).
mRNA delivery

Figure 9. Efficient RNA-based gene knockout in vitro. (A) IVT Cas9 mRNA was transfected into HEK293T-EGFP cells with two types of EGFP-targeting gRNA. EGFP expression in non-treated (NC) and transfected cells was observed by microscopy (B) and quantified using flow cytometry (C-D). (E-F) The editing to EGFP genes on the genome was further confirmed by T7E1 assay and Sanger sequencing.
Cas9-gRNA RNP delivery

Figure 10. CRISPR-mediated gene knockin in iPSCs. UBC-driven EGFP (2432 bp) was delivered into iPSCs by electroporation of Cas9/gRNA RNP complex and donor vector. Successful gene knockin was confirmed by (A) Sanger sequencing and (B) PCR genotyping of single clones (WT locus: 762 bp; EGFP: 3194 bp). (C) Representative fluorescence microscopy image. (D) Karyotyping results. (E) Representative immunofluorescence stainings of pluripotency markers NANOG, OCT4, and SOX2 in EGFP knockin iPSCs confirming maintained pluripotency.
Resources
FAQ
Either shRNA-mediated knockdown or nuclease-mediated knockout (e.g. CRISPR or TALEN) can be valuable experimental approach to study the loss-of-function effects of a gene of interest in cell culture. In order to decide which method is optimal for your specific application, there are a few things you should consider.
Mechanisms
Knockdown vectors
Knockdown vectors express short hairpin RNAs (shRNAs) that repress the function of target mRNAs within the cell by inducing their cleavage and repressing their translation. Therefore, shRNA knockdown vectors are not associated with any DNA level sequence change of the gene of interest.
Read more about our shRNA knockdown vectors
Knockout vectors
CRISPR and TALEN both function by directing nucleases to cut specific target sites in the genome. These cuts are then inefficiently repaired by the cellular machinery, resulting in permanent mutations, such as small insertions or deletions, at the sites of repair. A subset of these mutations will result in loss of function of the gene of interest due to frame-shifts, premature stop codons, etc. If two closely positioned cut sites in the genome (i.e. within several kb) are targeted simultaneously, this can also result in the deletion of the intervening region.
Read more about our CRISPR vectors
Effectiveness
shRNA-mediated knockdown will never completely repress the expression of the target gene. Even for the most effective shRNAs, some residual expression of the target gene will remain. In contrast, in a fraction of treated cells, CRISPR and TALEN can generate permanent mutations which may result in complete loss of gene function.
Consistency and uniformity
shRNA vectors generally provide high cell-to-cell uniformity within the pool of treated cells and very consistent results between experiments. In contrast, CRISPR and TALEN produce results that are highly non-uniform from cell to cell due to the stochastic nature of the mutations introduced. To fully knock out the gene of interest in a cell, all copies of the gene in the cell must be knocked out. Given that normal cells have two copies of any gene (except for X- or Y-linked genes) while cancer cells can have more than two copies, such full knockout cells may represent a very small fraction of all the treated cells. For this reason, nuclease-mediated knockout experiments require the screening of clones by sequencing to identify the subset in which all copies of the gene of interest have been knocked out.
Off-target effects
Off-target effects have been reported for both shRNA-mediated knockdown and nuclease-mediated knockout. The off-target phenotype(s) can be estimated by using multiple different shRNAs to target the same gene. If a gene knocked down by multiple different shRNAs results in consistent phenotype(s), then it argues against the phenotype(s) being caused by off-target effects. For CRISPR- or TALEN-mediated knockout, multiple clones containing loss-of-function mutations should be analyzed in order to account for any phenotype(s) that may be due to off-target mutations. Additionally, bioinformatically identified off-target sites could be sequenced in the clones to see if they have been mutated.
Both CRISPR and TALEN systems have been harnessed to edit genomes of cultured cells and model organisms. Both systems can be used to knock out genes, or to knock in point mutations or insertions, but these two systems are different in several ways and have their own pros and cons.
Mechanisms
CRISPR
The CRISPR system uses a site-specific guide RNA (gRNA) to direct the Cas9 nuclease to its target site in the genome to create DNA cleavage. The target sequence is typically ~20 bp long, and sites containing a few mismatches may still be recognized and cleaved.
Read more about our CRISPR vectors
TALEN
The TALEN system employs a pair of chimeric proteins, each composed of a TAL effector DNA-binding domain (recognizing a specific sequence) fused to a FokI nuclease domain. The pair of proteins are designed to bind to a pair of target sites in the genome, each ~18 bp long and flanking a 14-20 bp spacer. Upon binding to DNA, the Fokl nuclease domains on the pair of proteins are able to dimerize, which in turn leads to DNA cleavage within the spacer region between the two target sites.
Efficiency
Both CRISPR- and TALEN-mediated genome editing show good efficiency, but the efficiency varies a lot depending on application, species and cell type. In general, CRISPR can be delivered into cells and induce DNA cleavage more efficiently than TALEN.
Off-target effects
A CRISPR gRNA targets ~20 bp sequence, whereas a TALEN pair binds to a total of ~36 bp target sequence. In addition, Cas9/gRNA complex has higher tolerance for sequence mismatches (up to 5 bp mismatches) than TALEN does. Therefore, TALEN-mediated cleavage has better specificity than CRISPR, and off-target cleavage in the genome by TALEN is unlikely. In contrast, off-target effects have been reported for CRISPR in cell lines, though analyses of CRISPR knockout mice suggest lower off-target frequency in vivo. Recent developments of CRISPR system have significantly enhanced CRISPR specificity. By using Cas9 nickase (Cas9 mutant that contains only one catalytic nuclease domain, e.g. Cas9_D10A and Cas9_H840A) with dual gRNAs, two single-strand DNA nicks are generated with close proximity of the target region, resulting in a double-nick DSB (double-strand break) within the target region that could be repaired. In this design, the off-target effects are minimized since the dual gRNAs expand the target sequence to ~40 bp long.
Target site requirements
TALEN can be generated to specifically target nearly any sequence in the genome. In contrast, target site selection for CRISPR is limited by the requirement for a PAM sequence (typically NGG) sequence located on the immediate 3’ end of the gRNA target sequence. This is no barrier to knocking out genes because cleavage anywhere in the gene is potentially effective but may present difficulties in generating site-specific mutations or insertions that require cleavage at a specific position of the gene. To precisely edit a specific genomic site using CRISPR, a homologous recombination donor vector or long oligo containing the desired edit sequence flanking by the immediate upstream and downstream homology arms of the target site can be delivered to the cells together with gRNA(s) and Cas9, in order to guide HDR (homology directed repair)-mediated DNA repair at the target site.
Design your homologous recombination donor vector online
Technical challenges
In terms of technical simplicity, CRISPR out competes TALEN in several ways. First, for vector construction, CRISPR system only needs to construct a short gRNA because targeting of Cas9/gRNA complex relies on simple RNA/DNA hybridization, while TALEN system requires re-engineering of the TAL DNA-binding domain that is unique for each protein-DNA interaction. Therefore, gRNAs are cheaper and easier to design and construct than TALENs which always require two vectors per target site. Secondly, for some applications, such as injecting mouse embryos, Cas9 protein and gRNA can be more efficiently delivered via direct injection, but TALEN cannot. Thirdly, CRISPR is extremely versatile in genetic screening experiments since CRISPR screening library expressing many thousands different gRNAs can be easily constructed in a high-throughput manner.
For CRISPR-mediated genome editing, Cas9 nuclease is directed to the target site of site-specific guide RNA (gRNA) in the genome to create DNA cleavage. In most cases, to generate simple gene knockout, a single gRNA can be used together with Cas9 to generate a double-strand break (DSB), which is then inefficiently repaired by the non-homologous end joining (NHEJ), resulting in permanent mutations, such as small insertions or deletions, at the site of repair. A subset of these mutations will result in loss of function of the gene of interest due to frame-shifts, premature stop codons, etc.
Dual gRNAs can be used if Cas9(D10A) nickase is being used to target the two opposite strands of a single target site. In this approach, the nickase enzyme will generate single strand cuts on both strands, one guided by each of the two gRNAs, resulting in DSBs at the target site. Generally, this method reduces off-target effects of CRISPR/Cas9 expression because targeting by both gRNAs is necessary for DSBs to be generated.
Dual gRNAs can also be used when Cas9(D10A) nickase and an exogenous donor DNA template are being used to introduce specific base-changes (e.g. knockins) into a gene of interest. In this approach, the two opposite strands would be targeted by the two gRNAs at two sites flanking the desired mutation site, and homology-directed repair (HDR) pathways make use of the exogenous donor template to repair the excised sequence.


